
Abstract
The atomic‐scale structure of  solid solutions, previously experimentally determined by extended x‐ray absorption fine structure, is used to model the strain energy associated with the local distortions in bondlengths and bond angles relative to those in the pure binary compounds. The distortion energy associated with the bondlength change was calculated to be
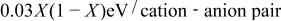 , where
 is the
 mole fraction, and adding the contribution from the bond‐angle distortions leads to a total strain energy of
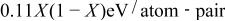 . The composition dependence of the strain energy is the same as the regular solution enthalpy of mixing,
 , and the calculated value of the solid solution interaction parameter,
 , is in good agreement with the reported experimental value of 0.13 eV/atom‐pair for
 . Similar calculations are made for four other zincblende solid solutions for which the atomic‐scale structure has been determined, and the results suggest that local bond strain is the major contribution to the enthalpy of mixing. Generalization of the strain calculations leads to a second‐order dependence of the
 on the normalized difference in endmember lattice constants, with no adjustable parameters, and this result is compared to semiempirical models for
 .
Access this article
The computer you are using is not registered by an institution with a subscription to this article. Please choose one of the options below.
Purchase from
Purchase this article from our trusted document delivery partners.

Make a recommendation
To gain access to this content, please complete the Recommendation Form and we will follow up with your librarian or Institution on your behalf.
For corporate researchers we can also follow up directly with your R&D manager, or the information management contact at your company. Institutional subscribers have access to the current volume, plus a 10-year back file (where available).
